Medical Disposable Manifolds Factory: Key Considerations for Quality and Compliance
2026-09-03
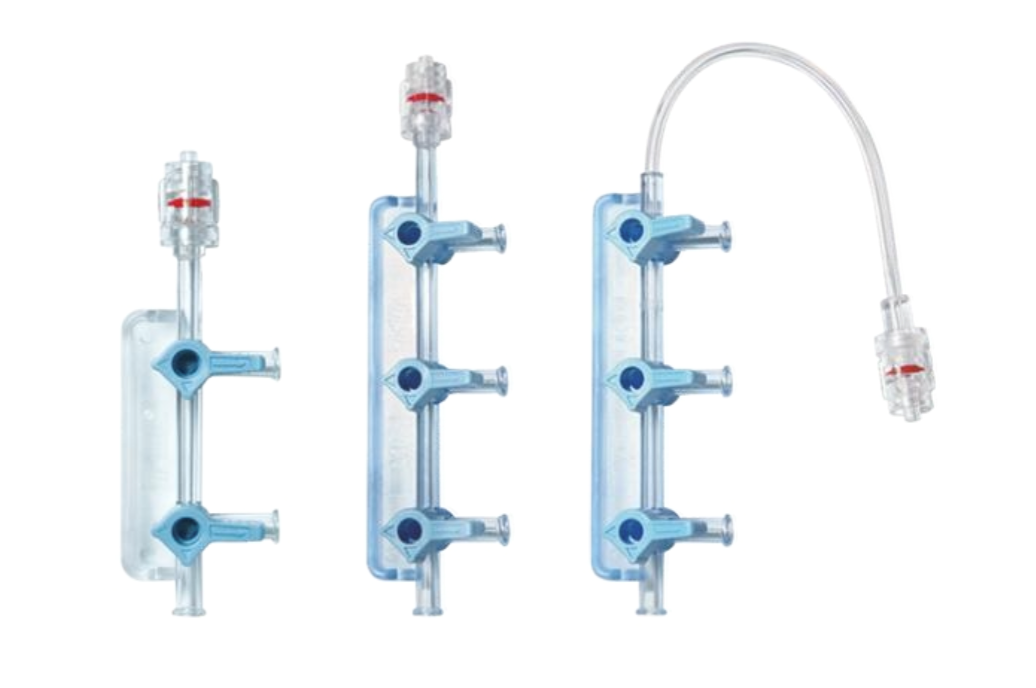
Choosing the right factory for medical disposable manifolds isn’t just a sourcing decision—it’s a direct line to patient safety and regulatory success. As these components sit at the heart of fluid management in countless clinical procedures, even small deviations in material quality or production consistency can trigger serious downstream failures. That’s why forward-thinking procurement teams are looking beyond price per unit and digging into the real indicators of reliability: cleanroom protocols, validation documents, traceability systems, and a supplier’s willingness to stand behind every lot. Among manufacturers that consistently clear these hurdles, INT has emerged as a name worth watching—not because of flashy marketing, but because their contamination control and compliance track record keeps showing up in audits you can actually verify. If you’re tired of vague claims and want a factory checklist that separates genuine quality from wishful thinking, the following guide is exactly what the doctor ordered.
Raw Material Decisions That Impact Every Batch
Every batch starts long before the first ingredient is measured. The choice of raw materials sets the baseline for consistency, flavor, and even how the final product behaves on the shelf. A supplier change, a subtle shift in moisture content, or a different growing region can ripple through the entire process without warning.
Small variations in raw inputs rarely announce themselves. One lot of flour might absorb water differently, forcing bakers to adjust mixing times on the fly. Another batch of spices could carry a slightly higher oil content, shifting the aroma profile just enough to be noticed by regular customers. These are not failures—they are the daily reality of working with natural materials.
The most resilient operations treat raw material evaluation as an ongoing conversation, not a one-time checklist. By tracking how each ingredient performs across multiple batches, teams build a working knowledge that no spec sheet can replace. That accumulated understanding is what separates a product that is merely acceptable from one that stays reliably excellent, batch after batch.
Cleanroom Protocols Beyond the Standard Checklist
Most cleanroom failures can be traced back to something a checklist never asks: how people move once they're inside. Gowning correctly is only the beginning; if a technician adjusts their goggles or touches a wall to steady themselves, they've already compromised the very environment they worked so hard to protect. A robust protocol goes beyond verifying glove and hood placement by training operators to treat every motion as a potential particle release. Slow, deliberate movements, avoiding cross-body reaches over open product, and never turning your back to a unidirectional airflow stream are habits that no static checklist can enforce. When we audit, we watch boot cover wear patterns, face mask adjustment frequency, and whether people instinctively lean against workbenches — these small tells reveal more about contamination risk than a signed gowning log.
Particle counts and pressure differentials are commonly logged, but the numbers themselves hide more than they reveal. A cleanroom can read within spec at 8:00 a.m. and still be dangerously unstable during an active material transfer at noon. True protocol extends to measuring how quickly the room recovers after a door opens, whether temperature spikes near heat-generating equipment create micro-eddies that trap particles, and whether pass-through chambers reach their intended pressure cascade before use. Smoke studies should not be a one-time certification exercise; they need to be repeated whenever equipment layouts change or new personnel are introduced. The goal isn't to confirm the room works in theory, but to know exactly where the airflow gets lazy under real operational load.
Cleaning checklists tend to focus on visible surfaces, but contamination rarely announces itself in plain sight. Disinfectant contact time, residue accumulation from repeated wiping, and rotation schedules to prevent microbial resistance are all too often reduced to a single line item: 'wipe down work area.' What matters more is what gets missed — the undersides of carts, castor wheels, transfer hatch gaskets, and the seams where panels meet. We've found ATP swab failures on telephones and intercom buttons even when bench tops passed with flying colors. A rigorous program rotates random sampling of these overlooked zones, treats repeated low-level hits as early warnings, and adjusts cleaning frequency based on data rather than habit. In the end, protocols that survive contact with reality are less about adding more checks and more about questioning the ones you already trust.
Sterilization Validation Without Trial and Error
Traditional sterilization validation often leans on iterative cycle development—adjusting temperature, time, or load configuration until a biological indicator is fully inactivated. This approach, while familiar, consumes production hours and introduces variability. A more deterministic path uses quantifiable microbial death kinetics. By establishing the D-value (decimal reduction time) and z-value of the reference organism under actual load conditions, one can calculate the required lethal dose directly. This shifts the process from guesswork to a calculated physical endpoint.
Physical qualification of the sterilizer supports this approach. Mapping temperature distribution and pressure profiles across worst-case load configurations identifies cold spots and air pockets without running repeated biological indicator cycles. Once the slowest-to-heat location is characterized, correlation with microbial resistance data allows prediction of the minimum F0 or equivalent lethality. The validation run then becomes a confirmation of a pre-established model, rather than a search for parameters.
The practical outcome is fewer aborted cycles and faster release. Instead of testing several candidate cycles against biological indicators, engineers set acceptance criteria from calculated lethality margins and use routine physical monitoring to verify continued compliance. This reduces reliance on retrospective pass/fail outcomes and supports a risk-based, science-driven validation lifecycle that adapts quickly to new products or load patterns.
Lot-Level Traceability From Pellet to Package
Manufacturing a single batch often starts with raw pellets arriving in bulk. Each container receives a unique lot identifier before it enters production, linking the supplier’s data to the internal workflow. That identifier stays with the material through every blending, sieving, or compaction step, so operators can pinpoint exactly which pellet lot contributed to any downstream unit.
By the time the final product is sealed into bags or bottles, each package carries a machine-readable code tied back to the original pellet lot. This isn’t just a theoretical link—line sensors and weigh stations log events automatically, capturing time, operator ID, and equipment status. If a defect appears in one carton, the traceability data narrows the investigation to a specific pellet delivery rather than an entire week of production.
Recalls become surgical instead of blanket. A packaging line supervisor can pull up the entire history of a single lot in minutes, seeing which silo held the pellets, how long they sat before processing, and which shift handled the conversion. That level of granularity also helps with supplier accountability—when a pellet batch underperforms, the data makes the conversation factual, not speculative.
Preparing for Regulatory Audits Without the Rush
Most teams treat audit preparation as a deadline-driven scramble, but the calmest responders build a habit of evidence hygiene long before a notice lands. Each control gets a single owner, a set review cadence, and a clear pointer to the documents that prove it works. When a regulator asks for the third-party access review from eight months ago, the answer is a file path, not a panicked search through email threads.
Run a quarterly 'friendly audit' where someone outside the compliance group requests the same artifacts an auditor would. Time the response, list what was missing or outdated, and then assign the gaps as normal engineering tasks. This turns vague anxiety into a backlog of small fixes—none of which feel urgent until they suddenly are.
Finally, coach the people who will sit in the room. A two-sentence explanation of why a control exists beats a perfect policy document that nobody can summarize. Brief walkthroughs on likely questions remove the freeze response and let the team show the judgment behind the process, which is what auditors actually want to assess.
Turning Production Data Into Continuous Quality Gains
Most factories already collect production data, but few treat it as a living feedback loop. The real shift happens when a rejected batch stops being a spreadsheet entry and starts triggering an immediate adjustment on the line. Instead of waiting for weekly quality reports, operators see drift in torque values or coating thickness the moment it occurs, and can correct the process before defects multiply. That kind of real-time visibility turns passive logging into an active quality tool.
Continuous gains come from linking data points that usually live in separate silos. For example, a spike in surface roughness might correlate with a specific supplier’s raw material lot, or with a tool change that happened two hours earlier. By joining machine parameters, environmental readings, and inspection results into one coherent story, teams can uncover root causes that never appeared in manual audits. Over time, these patterns become decision rules—not just alarms—that guide preventive maintenance and operator training.
The most valuable production data doesn’t just describe the past; it predicts the next likely failure. A simple trend in vibration frequency or cooling rate can signal that a process is drifting toward its control limit long before a bad part is produced. When this predictive layer is combined with small, daily experiments on the line, quality stops being an end-of-line gate and becomes a continuous, compounding improvement cycle. Each shift leaves the process slightly more stable than the previous one, driven by evidence rather than guesswork.
FAQ
Incoming polymer and component inspection, process validation for injection molding, in-process leak and dimensional checks, environmental monitoring of cleanrooms, post-sterilization sterility testing, and final package seal verification. Each of these directly affects product safety or functionality.
Manifolds contact patient fluids or respiratory gases, so particles or microbes introduced during molding or assembly can cause serious harm. Most factories use ISO Class 7 or 8 cleanrooms with controlled temperature, humidity, and pressure differentials. Operators wear sterile garments and follow strict gowning procedures; surfaces and air are routinely sampled for viable and non-viable particulates.
ISO 13485 is the baseline quality management system, with additional requirements from FDA 21 CFR Part 820 for US markets and EU MDR 2017/745 for Europe. Product-specific standards like ISO 10993 for biocompatibility and ISO 80369 for small-bore connectors often apply. Compliance means maintaining a technical file or design dossier and being ready for unannounced audits.
Medical-grade polymers such as polycarbonate, polypropylene, and COC are common. The material must withstand the intended sterilization method (ethylene oxide, gamma, or electron beam), resist stress cracking, maintain transparency if visual inspection is needed, and pass ISO 10993 biocompatibility tests. It should also be free of DEHP and other restricted substances.
Each lot receives a unique identifier that links raw material batches, molding machine parameters, assembly operators, QC inspection records, sterilization batch, and distribution records. Many factories use ERP systems with barcode scanning at each station. This allows a complete forward and backward trace in minutes if a recall or complaint arises.
Every production lot undergoes leak testing (pressure decay or vacuum), flow resistance measurement, burst pressure testing on samples, connector gauge checks, particulate count, and endotoxin testing. Sterile products require sterility testing per pharmacopeia methods, and package integrity is verified by dye penetration or seal strength tests.
A manifold often includes valves, caps, tubing, and bonded joints from multiple suppliers. A single substandard O-ring or connector can compromise the entire assembly. Factories typically audit suppliers for ISO 13485 compliance, perform incoming inspection on every lot, and require change notification for any material or process modification. This reduces variability before parts ever reach the production floor.
Flash around sealing surfaces, short shots, bubbles or voids in transparent parts, weak weld lines, blocked lumens from improper core pin alignment, mismatched connector dimensions, and surface scratches that harbor bacteria. Most defects can be prevented through proper mold maintenance, validated process parameters, and automated vision inspection.
Conclusion
In a medical disposable manifolds factory, the choices made at the raw material stage ripple through every downstream process. Resin selection isn't just about cost or moldability; it must align with biocompatibility expectations under ISO 10993 and withstand the aggressive solvents or drugs that the manifold may contact. Switching suppliers without a full equivalency study can alter extractables profiles or compromise bonding strength, so material locks and incoming lot audits become non-negotiable. On the production floor, cleanroom discipline goes far beyond a generic checklist. Operators must understand why fingertip plates are placed near critical zones, why air pressure differentials favor the filling area, and why garment protocols shift when moving from ISO 8 to ISO 7 spaces. Real compliance emerges when staff grasp the reasoning behind each rule, not just the procedure. Sterilization validation similarly demands a risk-based mindset. Instead of repeating a fixed cycle and hoping for the best, engineers map the worst-case bioburden, determine the required SAL, and confirm that the chosen dose or cycle delivers a reproducible lethality across the entire manifold lumen—especially in dead-leg areas where fluid can stagnate.
Traceability is equally critical, and it must extend from the plastic pellet to the final sealed pouch. Lot-level records that tie together the resin batch, extrusion parameters, assembly line, and sterilization load enable a rapid and precise recall if a defect ever surfaces. This same granular data stream becomes the backbone of audit readiness, eliminating the last-minute scramble for paper records. Instead of treating audits as an external threat, factories that maintain a live quality dossier and routinely run mock inspections can invite auditors into a well-organized system where nonconformances are already documented with corrective actions. Finally, production data should not sit idle in spreadsheets. By tracking yield, defect types, and environmental monitoring trends over time, a factory can spot subtle drifts—such as a gradual increase in particulate counts or a slight shift in seal strength—before they become batch failures. Turning that data into monthly cross-functional reviews closes the loop, making quality a continuous, measurable outcome rather than a static certificate.
Contact Us
Contact Person: Jeffrey
Email: [email protected]
Tel/WhatsApp: 86-0633-2230056
Website: https://www.sd-intmedical.com